Integrated preclinical discovery, screening, potency analytics, and full-spectrum safety studies validated under stringent GLP documentation.
As global health authorities elevate thresholds for safety margins and reproducible characterization, traditional contract toxicology is being transformed by multimodal testing matrices.
Unlike single-molecule small synthetics, botanical extracts and polyherbal formulations contain hundreds of secondary metabolites, volatile terpenes, and complex macromolecules. This intricate chemical landscape presents unique nonclinical toxicity challenges: xenobiotic synergy, organ-specific accumulation, potential CYP450 enzyme inhibition, and batch-to-batch variation.
Our ODM toxicity services bridge raw botanical screening with validated toxicology packages, turning empirical herbal knowledge into scientifically quantifiable, regulator-ready dossiers.
MedGence (Yineng Pharma) operates at the convergence of classic medical knowledge and modern toxicological verification. Having completed modern medical safety profiles for 83 classic Traditional Chinese Medicine (TCM) formulas, developed 22 innovative natural medicines, and registered 56 hospital formulations, our multidisciplinary team ensures every assay conforms to international testing guidelines.
Our research protocols adhere rigorously to OECD Good Laboratory Practice (GLP), ICH S-series safety guidelines, and national regulatory pharmacopeias.
Generic contract manufacturers provide standard pass/fail readouts. As an advanced ODM Toxicity Studies Manufacturer, we deliver deeper contextual insights: Target Organ Toxicity Identification (TOTI), Maximum Tolerated Dose (MTD), No Observed Adverse Effect Level (NOAEL), and Toxicokinetic (TK) exposure modeling.
This mechanistic understanding provides clients with robust safety margins that prevent attrition during clinical phases and regulatory submission reviews.
From rapid early-stage cytotoxicity screens to pivotal GLP-compliant chronic safety and carcinogenicity packages, our facility covers the entire preclinical toxicity continuum.
Establish baseline safety parameters, target-organ susceptibility, and acute mortality thresholds under OECD 420, 423, and 425 guidelines.
Longitudinal evaluation of cumulative physiological impacts across 14-day, 28-day, and 90-day exposure windows under OECD 407 and 408 standards.
Compliant with ICH S2(R1) regulatory guidelines to detect point mutations, chromosomal damage, and systemic mutagenic liability.
Systemic functional safety suites assessing vital physiological systems prior to first-in-human clinical trials.
Investigate compound effects on male and female fertility, embryofetal morphogenesis, and postnatal developmental milestones under ICH S5(R3).
Interfacing bioanalytical chemistry with exposure modeling to evaluate systemic compound circulation, metabolite emergence, and organ clearance.
As regulatory authorities harmonize evidentiary standards while advancing ethical alternatives, manufacturers must implement forward-compatible preclinical testing strategies.
The signing of the FDA Modernization Act 2.0 represents a landmark regulatory evolution, authorizing the use of qualified alternatives to animal testing, including cell-based microphysiological systems, organoids, and computational (in silico) modeling to support Investigational New Drug (IND) applications.
Our ODM facility embraces this transition through a hybrid tiered model: we deploy 2D/3D human cell cultures, organ-on-a-chip platforms, and in vitro cytotoxicity screens as rapid early filters to eliminate high-risk structures, reserving targeted in vivo models for confirmatory systemic endpoints.
For international market authorizations spanning the United States (FDA), European Union (EMA), United Kingdom (MHRA), Japan (PMDA), and China (NMPA), data integrity and study transparency are critical. Under the OECD MAD framework, safety data generated in compliance with OECD GLP principles is accepted across all member and adhering nations.
MedGence guarantees standard operating procedures (SOPs), QA audit oversight, temperature-controlled sample traceability, and comprehensive archival systems designed to withstand international multi-jurisdiction regulatory audits.
Regulatory authorities increasingly reject nonclinical dossiers for botanical active ingredients if the test substance is insufficiently fingerprinted. A toxicity profile is scientifically meaningless if the exact chemical composition of the tested batch cannot be mapped to the commercial formulation.
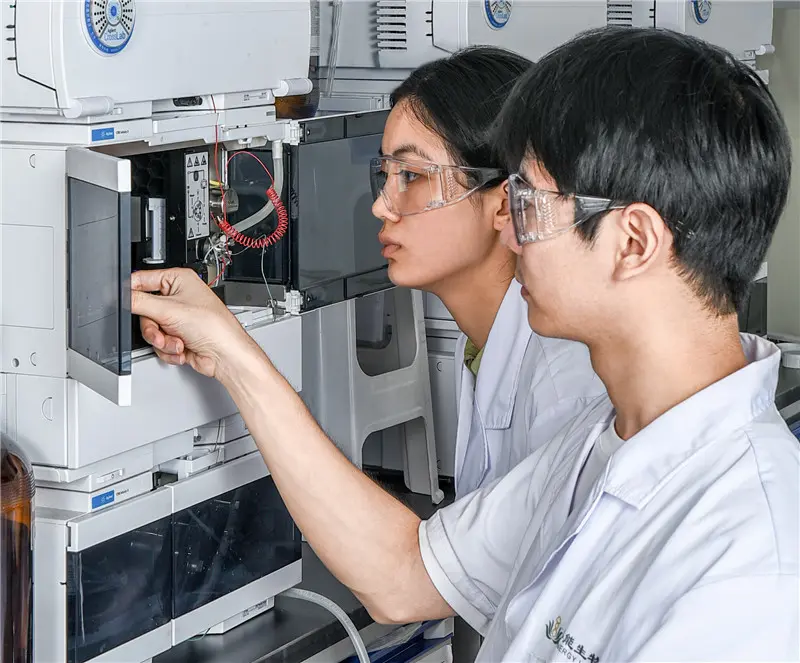
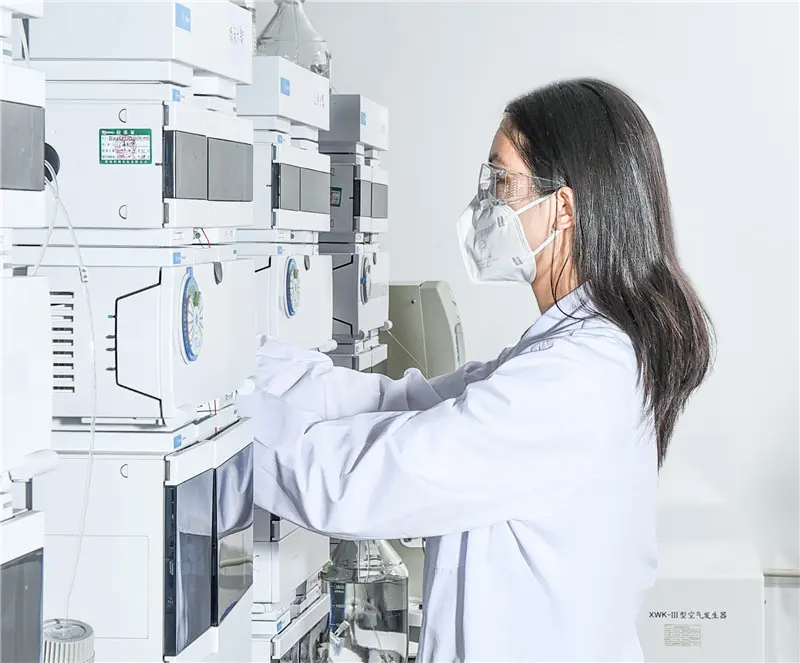
We solve this by coupling high-resolution chromatographic fingerprinting (HPTLC, HPLC-Q-TOF-MS), heavy metal residue quantification, and mycotoxin/pesticide screening directly with every toxicology batch, guaranteeing total compositional identity.
How our integrated CRO/CDMO campus delivers superior turnaround times, cost efficiency, and end-to-end quality control for international sponsors.
Western toxicology labs typically operate in isolation from botanical raw material supply chains, creating delays and chemical discrepancies. Based in Changsha, Hunan, MedGence leverages deep domestic access to raw botanical harvesting hubs, verified Good Agricultural and Collection Practices (GACP) farms, and dedicated extraction pilot plants.
Our extraction, purification, method development, and toxicology facilities exist under unified technical leadership, eliminating inter-vendor handoffs.
Global pharmaceutical and nutraceutical innovators face escalating R&D costs and multi-month queue times at major Western contract research labs. MedGence breaks this bottleneck by providing rapid study slot initiation within 2 to 4 weeks of protocol sign-off.
Our established nonclinical capacity and operational cost framework deliver 35% to 50% overall expenditure savings without compromising statistical rigor, GLP adherence, or analytical sensitivity.
Having authored monographs, extraction SOPs, and analytical quality standards for nearly 400 single-herb medicines and polyherbal preparations, our scientific bench is trained to anticipate matrix interference issues during in vitro and in vivo dosing.
This specialized domain expertise minimizes repeat assays, avoids dosage formulation precipitation, and accelerates regulatory submission pathways.
Tailored toxicology programs mapped to the distinct regulatory pathways of target markets and end-use product categories.
Structured nonclinical packages designed to satisfy FDA 505(b)(2), EMA Herbal Medicinal Product (HMPC), or NMPA Class 1/Class 2 traditional Chinese medicine regulatory filings.
Toxicological substantiation for New Dietary Ingredients (NDI), GRAS notices in the USA, and Novel Food approvals under EFSA regulations.
Cruelty-free, animal-free in vitro toxicology services aligned with the EU Cosmetic Regulation (EC No 1223/2009) and China CSAR safety dossiers.
Translating clinical hospital preparations into commercial registered drug products with modern nonclinical safety profiles.
Pioneering the integration of artificial intelligence, high-content screening, and microphysiological biochips to elevate safety prediction accuracy.
Standardized OECD/ICH animal models integrated with automated clinical pathology, targeted toxicogenomics, and LC-MS/MS bioanalysis to monitor metabolic cascades and tissue changes.
Automated fluorescence microscopy to simultaneously assess mitochondrial membrane potential, reactive oxygen species (ROS) induction, and apoptosis across diverse cell lines.
Microfluidic Liver-on-a-Chip and Gut-Kidney vascularized axes that mimic physiological mechanical shear stresses, providing accurate human-relevant organ toxicity data.
Machine-learning models trained on proprietary phytochemical datasets to predict off-target bioactivity, mutagenicity, and cardiotoxicity before wet-lab synthesis or extraction.
How our quality assurance framework shields sponsors from regulatory discrepancies, inspection non-conformities, and intellectual property hazards.
Our quality management division operates independently of laboratory study directors, reporting directly to the executive board. Every study phase undergoes three-tier protocol auditing: protocol review, in-life phase inspection, and final raw data/report cross-verification.
Laboratory data capture systems comply with US FDA 21 CFR Part 11 electronic records, complete with automated audit trails, user privilege stratification, and immutable cloud-synced backups.
Since 2016, the FDA requires nonclinical study data for IND, NDA, and BLA submissions to be formatted in the Standard for Exchange of Nonclinical Data (SEND). Non-SEND files encounter immediate refusal-to-file (RTF) risks.
Our computational bio-statistics unit maps all in-life observations, laboratory findings, and histopathological terminology into CDISC-compliant SEND datasets, ensuring effortless eCTD module 4 submissions.
As an ODM/CDMO partner, our business model relies entirely on contract research; we do not commercialize competing retail brands. All formulas, purification methods, extraction parameters, and study data remain 100% client-owned intellectual property.
Sponsors receive secure project firewalls, dedicated cleanroom pilot manufacturing suites, and non-disclosure governance backed by international legal jurisdiction.
Clear answers for toxicologists, regulatory directors, and procurement leaders evaluating nonclinical study programs.
Seamlessly extend your nonclinical pipeline from safety characterization to clinical trial validation and scalable finished dosage production.
Consult directly with our toxicologists and regulatory directors to scope your toxicology package, confirm turnaround timelines, and receive itemized budgetary plans.