Accelerating translational phytochemical research through vertically integrated development workflows, robust analytical chemistry, and regulatory-ready documentation.
Transitioning from empirical extraction to precision molecular phytotherapy under rigorous cGMP and ICH standards.
Global regulatory bodies including the US FDA (Botanical Drug Guidance), EMA (HMPC monographs), and NMPA mandate reproducible multi-target molecular characterization. Pure crude extracts no longer suffice without comprehensive chromatographic fingerprinting and batch traceability.
Global pharmaceutical and nutraceutical innovators are increasingly divesting physical synthesis infrastructure. Entrusting end-to-end development to specialized CRO/CDMO platforms reduces discovery overhead by up to 42% and accelerates regulatory time-to-market.
The geographical variation of botanical raw materials creates acute supply chain risks. CDMOs equipped with multi-origin phytochemical evaluation and sustainable agro-tech partnerships offer critical resilience against harvest vulnerabilities and active ingredient fluctuations.
Combining 14+ years of proprietary natural chemistry with over 300 in-house research scientists, serving global tier-1 pharmaceutical groups.
A closed-loop industrialization paradigm engineered to eliminate attrition risks at every translational inflection point.
High-content phenotypic screening, multi-target affinity selection, and database-driven botanical bio-fractionation across alkaloids, saponins, and polysaccharides.
Establishing ICH-compliant qualitative and quantitative assays using high-performance thin-layer chromatography (HPTLC), HPLC-MS/MS, and GC fingerprint profiles.
Comprehensive GLP-compliant in vitro bioassays, in vivo murine disease models, LD50 acute toxicity, chronic sub-acute exposure, and full genotoxicity panels.
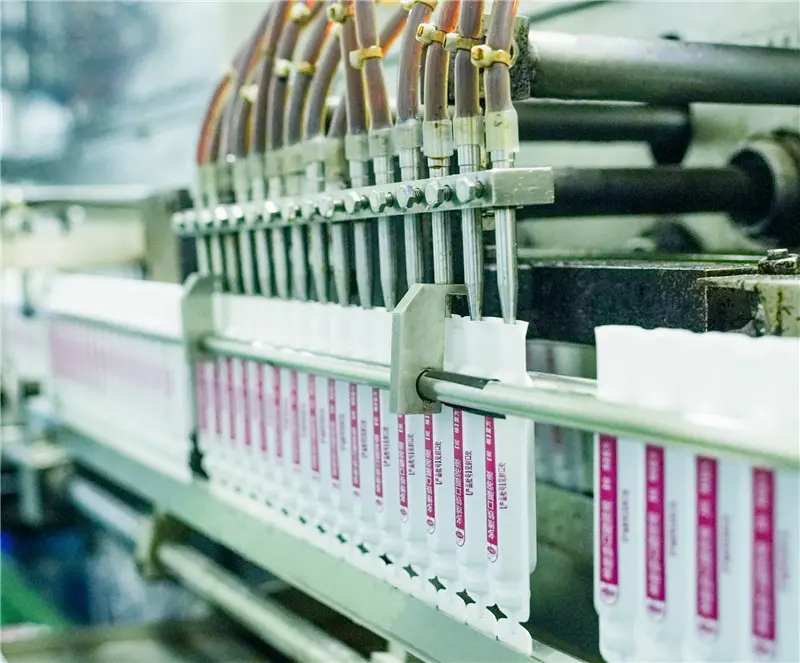
Seamless tech transfer to GMP pilot plants, Phase I–IV clinical batches, and high-volume commercial finished dosage manufacturing across multiple formats.
Formulating compliant, scientifically validated functional matrices tailored to distinct regional markets and regulatory frameworks.
Structured chemistry, manufacturing, and control (CMC) dossiers supporting botanical Investigational New Drug (IND) and New Drug Applications (NDA) for FDA, EMA, and NMPA submission.
Standardized active principles supported by human micro-trials and biological mechanistic data, enabling defensible health claims across North American, European, and Asia-Pacific retail spaces.
Standardized phytonutrient complexes with in vitro validated dermal barrier restoration, anti-inflammatory pathways, and photo-aging protection properties compliant with INCI mandates.
Clear, technical, and regulatory guidance for procurement managers, R&D directors, and chief scientific officers.
Traditional toll processors merely execute client recipes without chemical insight. A full-scope CDMO like MedGence engineers the underlying extraction science, develops and validates analytical HPLC/HPTLC protocols, optimizes bioactive yield, performs GLP safety testing, and ensures regulatory compliance across all deliverables.
We systematically map botanical species across geographic coordinates, microclimates, and harvest periods. By correlating these environmental factors with HPLC active compound profiles, we establish strict sourcing criteria and formulation adjustments that guarantee commercial batch-to-batch consistency.
We work exclusively under comprehensive Non-Disclosure and IP Assignment Agreements. All chromatographic libraries, extraction kinetics, purification parameters, and clinical documentation generated during contract research become the exclusive intellectual property of the commissioning client.
Our collaborative and in-house facilities manufacture concentrated liquids, vacuum-dried extract powders, pastes, volatile essential oils, aqueous oral liquids, hard-gelatin capsules, softgels, enteric-coated tablets, and multi-component soluble effervescent granules.
Yes. Beyond preclinical cell and rodent bio-assays, our team structures and executes GCP-compliant Phase I through Phase IV clinical studies. We manage patient cohorts, monitor biomarker endpoints, and compile complete clinical documentation ready for regulatory drug filings.
Feasibility assessment and raw material fingerprinting typically require 2 to 4 weeks. Full method establishment, extraction scale-up, and preclinical toxicology panels average 8 to 16 weeks, followed by commercial tech-transfer to our pilot and large-scale manufacturing facilities.
Explore our specialized production lines and certified testing facilities engineered for international volume requirements.