Direct contract access to specialized Phase I–IV clinical execution, validated analytical instrumentation, and commercial phytochemical isolation protocols.
MedGence (Yineng Pharma), headquarted in Changsha, Hunan, operates at the intersection of classical pharmacological heritage and modern molecular medicine.
In modern clinical development, post-marketing authorization is no longer the conclusion of therapeutic inquiry; rather, it marks the initiation of rigorous real-world therapeutic confirmation. Phase IV clinical trials represent the definitive operational benchmark for evaluating long-term safety, low-incidence adverse drug reactions (ADRs), therapeutic efficacy across non-homogeneous patient demographics, and health economics outcomes research (HEOR). For worldwide biopharmaceutical and botanical drug innovators, sourcing a high-caliber Chinese contract development and manufacturing organization (CDMO) and contract research organization (CRO) for Phase IV deliverables unlocks an unmatched convergence of clinical scale, deep pharmacokinetic profiling, and optimized manufacturing cost metrics.
MedGence (Yineng Pharma) operates at this vital nexus. Over 14+ years, our organization has executed clinical and technical transfer workflows for more than 100 industrial manufacturers, completed modern medical characterization of 83 classic Traditional Chinese Medicine (TCM) formulations, developed 22 novel proprietary natural medicine compounds, secured 56 national hospital preparation registrations, and institutionalized full-spectrum analytical release standards for approximately 400 single-herb active ingredients. When multinational buyers search for "China Phase Iv Manufacturers & Supplier," they require more than an assembly line; they demand an authoritative scientific partner capable of defending chromatographic repeatability, patient data integrity, and compliance across ICH, NMPA, US FDA, and EMA regulatory frameworks.
Phase IV trials administered within our Chinese facilities adhere strictly to ICH E6(R2/R3) Good Clinical Practice guidelines and NMPA Good Pharmacovigilance Practice (GVP) mandates. Data structures are constructed natively to export into electronic Common Technical Document (eCTD) Module 5 clinical study reports, guaranteeing cross-border acceptance by the US FDA and European Medicines Agency (EMA).
How domestic botanical agricultural access, biostatistical depth, and advanced physical pilot facilities eliminate manufacturing bottlenecks for commercial contract batches.
Our direct access to authentic "Daodi" geographical harvest regions across China eliminates primary material adulteration. We systematically quantify active chemical variability across geographic origins, altitudinal shifts, and multi-seasonal harvest windows to ensure steady phytochemical lot uniformity.
Bridging benchtop chemistry to multi-ton extraction. MedGence maintains in-house pilot plants coupled with partner cGMP manufacturing facilities directly supervised by our senior engineering cadres, preventing tech-transfer failures during scale escalation.
Client formulas, customized solvent fractionations, and proprietary manufacturing parameters are protected through entrusted contract research protocols. All operations operate under robust non-disclosure architectures with physical and digital isolation.
The primary barrier to successful Phase IV commercialization of natural products and botanical active pharmaceutical ingredients (APIs) lies in batch-to-batch chemical heterogeneity. Synthetic single-molecule pharmaceuticals maintain predictable crystallization profiles; conversely, complex plant extracts contain a wide matrix of primary metabolites, secondary polyphenols, flavonoids, saponins, and glycosides that oscillate based on rainfall, soil microbiome, and thermal extraction dynamics.
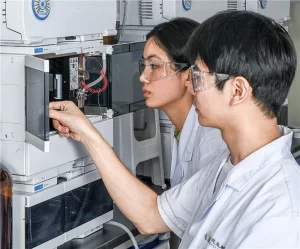
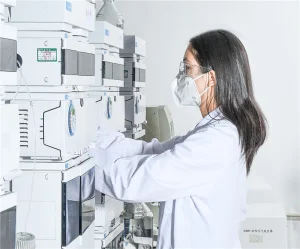
MedGence resolves this fundamental industrial challenge through multi-dimensional analytical fingerprinting. Before any Phase IV clinical trial or OEM manufacturing lot is released, raw botanical lots undergo High-Performance Thin-Layer Chromatography (HPTLC) chemical identification, High-Performance Liquid Chromatography (HPLC) active quantification, and Gas Chromatography (GC) volatile oil characterization. By locking in both specific marker ingredients and global chromatographic similarity thresholds (>0.95 similarity indices against verified reference standards), our manufacturing plants guarantee that post-marketing clinical trials in human populations correlate directly with documented chemistry, fulfilling the strictest evidentiary standards demanded by overseas buyers.
Bridging Chinese operational velocity with FDA 21 CFR Part 314.80, EMA Pharmacovigilance, and ICH GCP quality control.
Our clinical operational centers utilize validated, 21 CFR Part 11-compliant EDC platforms that permit real-time multi-center clinical record tracking. Clinical research associates (CRAs) conduct continuous source data verification (SDV) across participating hospital centers, ensuring clean statistical datasets ready for biostatistical aggregation.
With clinical trial registries expanding rapidly in Asia, localized Chinese hospital partnerships provide unmatched patient recruitment acceleration, enrolling targeted patient cohorts in months rather than years.
Post-market surveillance demands systematic adverse event (AE) reporting, Periodic Safety Update Reports (PSUR), and Risk Management Plans (RMP). MedGence provides end-to-end pharmacovigilance oversight, tracking rare drug-drug interactions, long-term hepatobiliary and renal markers, and demographic variations.
Our medical monitors maintain direct conduits with top-tier Class 3A clinical research centers, offering multinational brands complete regulatory alignment across both NMPA inspection standards and global health authority post-marketing commitments.
One of the most complex pain points for foreign biopharma procurement officers is whether Phase IV real-world observational data generated in China can be successfully utilized for foreign regulatory filings—such as US FDA Botanical Drug NDAs (under the FDA Guidance for Industry: Botanical Drug Development) or European Medicines Agency (EMA) Well-Established Use / Traditional Herbal Medicinal Products (THMPD) directives.
The solution engineered by MedGence is rooted in standardized harmonization. Because our Phase IV clinical methodologies incorporate CDISC (Clinical Data Interchange Standards Consortium) compliant SDTM (Study Data Tabulation Model) and ADaM (Analysis Data Model) data formats, our clinical study outputs can be ingested into international regulatory review dossiers without re-coding. Combined with accredited GLP preclinical toxicology validation (including LD50, Ames tests, and in vivo Micronucleus assays) and validated batch release records under ISO 17025 testing parameters, our clients obtain defensible scientific dossiers that overcome cross-border regulatory skepticism.
Aligning strategic enterprise procurement needs with disruptive industry advances in decentralized trials, AI safety monitoring, and dosage form innovation.
Phase IV trials are rapidly pivoting from small randomized samples toward prospective registry-based observational studies. By capturing Real-World Data (RWD) across diverse demographic strata, pharmaceutical sponsors confirm clinical effectiveness in real medical practice.
Integrating natural language processing (NLP) and machine learning models allows algorithmic analysis of millions of electronic health records (EHRs) to flag unexpected clinical events or subtle drug synergy patterns across patient sub-cohorts.
From highly hygroscopic botanical powders to uniform softgel capsules, oral liquids, and enteric tablets. Process chemistry guarantees stability, rapid dissolution kinetics, and bioavailable active payload integrity in finished goods.
Pharmaceutical procurement teams face intense corporate mandates: decrease clinical cycle times, mitigate supply chain volatility, ensure 100% trace-to-farm supply chain transparency, and reduce the Total Cost of Ownership (TCO) without exposing the parent company to regulatory scrutiny. When procuring Phase IV CDMO and clinical trial services in China, leading enterprises avoid fragmented transactional suppliers in favor of vertically integrated partners.
A unified CRO/CDMO partner like MedGence eliminates the classic "vendor friction" that destroys project profitability. Traditionally, biopharma brands hire one agency for raw material procurement, another laboratory for method development, a third for preclinical toxicology, a separate CRO for hospital clinical execution, and a distinct manufacturing facility for commercial tablet packaging. Every handoff introduces sample degradation risks, methodology translation errors, and IP leakage vulnerabilities. By centralizing these 12 interdependent disciplines under one scientific leadership team in Changsha, MedGence provides end-to-end accountability, reducing phase duration by 30–45% and eliminating redundant testing expenses.
How global enterprises deploy our Phase IV research, analytical testing, and CDMO infrastructure across actual product life-cycle stages.
Context: A multinational pharmaceutical enterprise secures market entry for an evidence-backed cardiometabolic botanical drug formulation, but health authorities condition approval upon a prospective 5,000-patient Phase IV observational safety registry monitoring hepatic and renal parameters over 24 months.
MedGence Execution: We mobilize protocol development across 25 collaborating Class 3A TCM and general university hospitals. Our data management teams configure cloud-based EDC with built-in audit trails, while our centralized bioanalytical core lab processes high-throughput serum metabolomics, generating clean interim and final safety dossiers that fulfill all regulatory commitments ahead of deadline.
Context: A commercial botanical formulation indicates strong clinical efficacy in adult metabolic syndrome, but the sponsor seeks to expand official labeling to include elderly populations (ages 65+) and adjust dosing for mild hepatic impairment.
MedGence Execution: Our clinical pharmacokineticists design a Phase IV open-label dose-titration protocol. Concurrently, our formulations laboratory establishes high-sensitivity LC-MS/MS bioanalytical assays to track key botanical marker metabolites in plasma, demonstrating pharmacokinetic bioequivalence and safety, allowing the client to execute a successful label expansion amendment.
Context: Sponsoring a comparative Phase IV study to prove that a modernized natural medicine plus standard-of-care protocol delivers superior cost-to-cure ratios and lower relapse rates than standard monotherapy alone.
MedGence Execution: We structure pragmatic, real-world comparative trials measuring Quality of Life (QoL) indices, total hospital days, and therapeutic recurrence rates. Statistical outcomes provide national health insurance reimbursement committees with empirical evidence supporting favorable formulary inclusion.
Context: An international nutraceutical brand holding a high-performing supplement seeks to elevate the product into an approved over-the-counter (OTC) botanical pharmaceutical by demonstrating clinical safety and rigorous batch standardization.
MedGence Execution: We execute standardized chemical fingerprinting, establish acute and chronic GLP toxicity profiles in animal models, and execute human Phase IV observational trials. Simultaneously, our CDMO facilities transition the formulation into standardized oral solid dosage forms manufactured under strict pharmaceutical GMP.
Our research programs and clinical trials interface directly with prestigious pharmaceutical manufacturers and premier university medical hospitals in China.
Detailed technical answers addressing regulatory compliance, bioanalytical validation, IP security, and contract delivery schedules.
Explore our core technological services and submit sample or RFP specifications directly to our scientific laboratory teams.